
> Basic Biology Concepts > DNA and Genetic Information Basics > Unraveling Simple Genetic Differences Across Populations
Unraveling Simple Genetic Differences Across Populations
We stand at the precipice of understanding life itself, a journey intricately linked to deciphering the subtle yet profound variations encoded within our DNA. This article embarks on a meticulous exploration of simple genetic differences that distinguish populations, unveiling how miniscule alterations at the molecular level forge the tapestry of human and species diversity. From geographical adaptation to disease susceptibility, these variations are not mere anomalies but powerful drivers of evolution and markers of ancestry.
Let us forge a surgical understanding of these distinctions, delving into concrete examples that illustrate their fundamental impact. We are initiating a thorough analysis here, armed with an expert perspective, to equip you with a strategic vision on how these differences shape phenotypes and evolutionary trajectories. Prepare to optimize your knowledge of the genetic forces that animate life, as mastering these concepts is key to understanding the fundamental principles of genetic material that orchestrate life. We will guide you through the mechanisms, implications, and practical applications of these variations, thereby transforming your perspective on population genetic dynamics. Each revelation is a biological opportunity unearthed, shaping a futuristic, accurate, and exhilarating understanding of biology.
Understanding Single Nucleotide Polymorphisms (SNPs) as Core Differences
We commence our exploration by dissecting Single Nucleotide Polymorphisms (SNPs), the bedrock of simple genetic variation. A SNP represents a variation at a single position in a DNA sequence, where one nucleotide is replaced by another. Imagine the vast genomic instruction manual; SNPs are like single typographical errors or deliberate substitutions occurring at specific pages. Globally, the human genome harbors approximately 10 to 30 million SNPs, with an average of one SNP every 100-300 base pairs. These variations are not random; they arise from replication errors, repair failures, or mutagenic exposures, and are subsequently passed down through generations, making them incredibly stable markers.
We distinguish between various types of SNPs based on their location and effect. Those found in coding regions (exons) can be synonymous (silent, not altering the amino acid sequence) or non-synonymous (missense or nonsense, leading to amino acid changes or premature stop codons, respectively). SNPs in non-coding regions (introns, intergenic) can still exert significant influence by affecting gene regulation, transcription factor binding, or mRNA stability. Our mastery over SNP detection, primarily through high-throughput sequencing and genotyping arrays, allows us to conduct powerful Genome-Wide Association Studies (GWAS). These studies correlate specific SNPs with complex traits or disease susceptibilities, such as diabetes, heart disease, or responses to medication. Understanding the distribution and functional impact of SNPs across populations is a crucial step in unraveling the genetic architecture of human diversity and disease.
Exploring Copy Number Variations (CNVs) and Their Population Impact
Beyond the single-base alterations of SNPs, we encounter Copy Number Variations (CNVs), which represent larger-scale structural differences in the genome. CNVs involve the duplication or deletion of segments of DNA, ranging from a few kilobases to several megabases. Picture a genomic blueprint where entire paragraphs or even chapters might be missing or duplicated; this is the essence of a CNV. These variations are generated primarily through mechanisms like unequal crossing-over during meiosis or non-allelic homologous recombination, processes that can rearrange significant portions of the genetic code.
The impact of CNVs on an organism's phenotype is often more pronounced than that of SNPs, owing to their size and the potential to alter gene dosage. A deletion might lead to a loss of function for specific genes, while a duplication could result in overexpression, both having critical biological consequences. We observe compelling examples of CNVs driving adaptive evolution; for instance, populations with diets rich in starch, like those in agricultural societies, often exhibit an increased number of copies of the amylase gene (AMY1), enabling more efficient starch digestion. Conversely, certain CNVs are strongly implicated in complex disorders such as autism spectrum disorders, schizophrenia, and Crohn's disease. Analyzing the frequency and distribution of CNVs across diverse populations provides us with a robust framework for understanding differential disease susceptibility, evolutionary pressures, and the intricate dance of genomic structural variation shaping life.
Mitochondrial DNA and Y-Chromosome Haplotypes: Tracing Ancestry
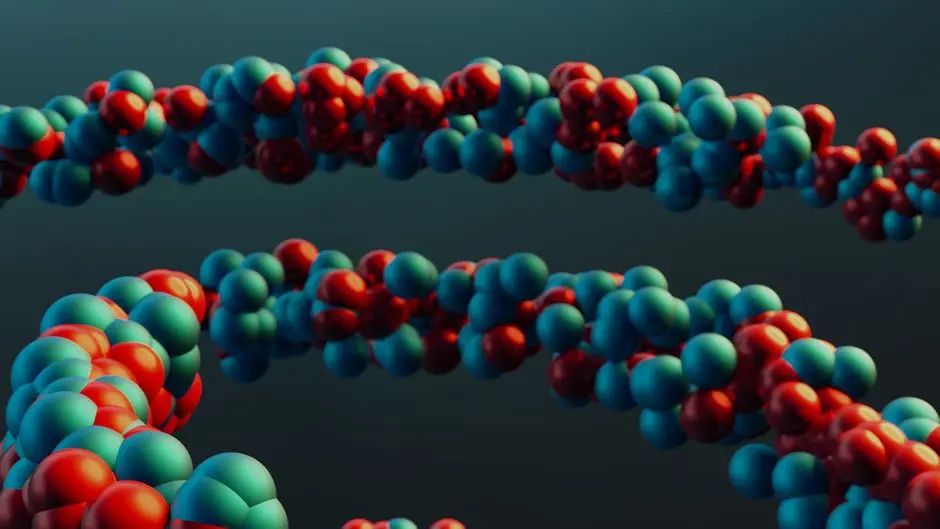
To trace deep ancestral lineages, we turn our focus to two unique genomic compartments: Mitochondrial DNA (mtDNA) and the Y-chromosome. These elements offer simplified, yet powerful, windows into population history due to their distinct modes of inheritance and relative lack of recombination. Mitochondrial DNA, found in the cell's powerhouses, is exclusively inherited from the mother. Its small, circular genome mutates at a faster rate than nuclear DNA, yet key regions remain highly conserved, making it an exceptional tool for tracking maternal lineages across millennia. The absence of recombination means that an individual's mtDNA sequence is a direct copy of their mother's, and her mother's, and so on, back through time.
Similarly, the Y-chromosome, present only in males, is passed directly from father to son. Apart from a small pseudoautosomal region, the majority of the Y-chromosome does not recombine with the X-chromosome, allowing paternal lineages to be traced with remarkable clarity. Scientists identify specific sets of genetic markers (SNPs or Short Tandem Repeats – STRs) on mtDNA and the Y-chromosome, which define distinct haplogroups. These haplogroups act as genetic signatures, revealing ancient migration routes, population bottlenecks, and the historical relationships between diverse human groups. For example, specific Y-chromosome haplogroups are prevalent in particular geographic regions, charting the dispersal of human populations out of Africa and across continents. By deploying these simple, non-recombining markers, we unlock unparalleled insights into the intricate tapestry of human genetic history and population dynamics.
Lactase Persistence and Sickle Cell Anemia: Case Studies in Adaptation
We consolidate our understanding of simple genetic differences by examining two iconic examples of human adaptation: lactase persistence and the genetic basis of sickle cell anemia. These cases vividly illustrate how single-gene variations, under specific environmental pressures, can confer significant selective advantages and become prevalent within populations. Lactase persistence, the ability for adults to digest lactose, is predominantly driven by a single SNP located upstream of the LCT gene. In most mammals, lactase production declines after weaning. However, in human populations with a long history of dairy farming (e.g., Northern Europeans, some African groups), this SNP maintains lactase expression throughout adulthood. This genetic difference provides a clear survival advantage in environments where dairy products became a crucial nutritional resource, demonstrating the power of positive selection acting on a simple genetic switch.
Similarly, sickle cell anemia, a severe genetic blood disorder, is caused by a single point mutation (Glu6Val) in the beta-globin gene (HBB). Individuals homozygous for this mutation develop severe anemia and other complications. Yet, the allele persists at high frequencies in malaria-endemic regions, particularly in parts of Africa, the Mediterranean, and South Asia. Why? Because individuals heterozygous for the sickle cell trait exhibit increased resistance to malaria, a deadly parasitic disease. This phenomenon, known as balancing selection, maintains the allele in the population, as the selective advantage against malaria outweighs the disadvantage of carrying one copy of the disease allele. These examples underscore how simple genetic differences are not just random variations, but often echoes of a population's historical interaction with its environment, shaping our very biology in profound ways.
Navigating the Future: Leveraging Simple Genetic Differences for Health and Conservation
Our journey culminates in a strategic outlook on leveraging these simple genetic differences to forge a healthier future and champion biological conservation. The insights gleaned from SNPs, CNVs, and haplogroups are not merely academic; they are actionable intelligence transforming fields from personalized medicine to ecological management. In pharmacogenomics, we harness individual SNP profiles to predict drug response and adverse effects, tailoring treatments to the patient's unique genetic makeup. For instance, specific variations in genes like CYP2D6 or VKORC1 guide clinicians in prescribing optimal dosages for antidepressants or anticoagulants, respectively, minimizing trial-and-error and maximizing efficacy. This empowers a proactive, precise approach to healthcare, moving beyond one-size-fits-all treatments to biologically optimized interventions.
Furthermore, these simple genetic markers are indispensable tools in conservation biology. By analyzing genetic diversity within endangered populations, we can assess their resilience, identify inbreeding risks, and formulate targeted breeding programs to ensure species survival. Forensic science also deploys these techniques, utilizing population-specific frequencies of genetic markers for identification and ancestry estimation. However, with this power comes responsibility. We must navigate the ethical landscape surrounding genetic privacy, potential discrimination, and informed consent. Our collective mission is to strategically deploy this genomic knowledge, ensuring its application is equitable, beneficial, and respects the profound diversity encoded within life's blueprint. We stand ready to optimize humanity's interaction with its genetic destiny, moving towards a future where every biological opportunity is meticulously seized.
Key Takeaways
SNPs (Single Nucleotide Polymorphisms)
These are single base-pair variations in DNA, foundational to genetic diversity. They are abundant in the human genome and influence everything from physical traits to disease susceptibility, making them key markers in genetic studies. Their location (coding vs. non-coding) dictates their functional impact.
CNVs (Copy Number Variations)
Representing larger DNA segment duplications or deletions, CNVs have a more significant impact than SNPs by altering gene dosage. They are crucial for understanding adaptation, such as increased amylase copies in starchy-diet populations, and are implicated in complex diseases like autism and schizophrenia.
Maternal and Paternal Tracers
Mitochondrial DNA (mtDNA) and the Y-chromosome are invaluable for tracing deep ancestry. MtDNA is maternally inherited without recombination, while the Y-chromosome is paternally inherited and largely non-recombining. Both form stable haplogroups, providing clear genetic signatures for human migration patterns and population histories.
Adaptation Examples
Lactase persistence (a single SNP) and sickle cell anemia (a single point mutation) are prime examples of simple genetic differences conferring evolutionary advantages. Lactase persistence benefits populations with dairy-rich diets, while the sickle cell trait offers malaria resistance in endemic regions, showcasing how natural selection operates on genetic variation.
Future Applications
Understanding simple genetic differences drives advancements in pharmacogenomics, allowing personalized medicine based on individual genetic profiles. It also plays a vital role in conservation biology for assessing population health and in forensics. Ethical considerations, however, must guide the responsible application of this powerful genomic knowledge.
FAQ
-
What is the primary difference between an SNP and a CNV?
The primary distinction lies in scale: an SNP (Single Nucleotide Polymorphism) involves a variation at a single base pair within the DNA sequence, such as a 'C' being replaced by a 'T'. In contrast, a CNV (Copy Number Variation) refers to the duplication or deletion of much larger segments of DNA, ranging from thousands to millions of base pairs. While SNPs are single-point changes, CNVs are structural rearrangements affecting significant chunks of the genome, potentially impacting multiple genes.
-
How do simple genetic differences contribute to disease susceptibility?
Simple genetic differences contribute to disease susceptibility by various mechanisms. An SNP in a coding region can alter a protein's function, or one in a regulatory region can affect gene expression levels. CNVs can lead to gene dosage imbalances, where too many or too few copies of a gene impair cellular processes. For example, specific SNPs are linked to increased risk for common complex diseases like type 2 diabetes, while certain CNVs are strongly associated with neurodevelopmental disorders. These variations don't guarantee disease but modulate an individual's predisposition when interacting with environmental factors.
-
Why are Mitochondrial DNA and Y-chromosome so useful for tracing ancestry?
Mitochondrial DNA (mtDNA) and the Y-chromosome are exceptionally useful for tracing ancestry due to their unique inheritance patterns and minimal recombination. MtDNA is inherited exclusively from the mother, meaning an individual's mtDNA sequence directly reflects their maternal lineage. The Y-chromosome, found only in males, is passed directly from father to son, thus tracing paternal lines. Crucially, the non-recombining nature of most of the Y-chromosome and the entirety of mtDNA ensures that mutations accumulate over generations without shuffling, creating distinct genetic 'haplogroups' that serve as stable markers of ancient human migrations and population relationships.
-
Can simple genetic differences lead to evolutionary advantages?
Absolutely. Simple genetic differences are often the very raw material upon which natural selection acts, leading to evolutionary advantages. A classic example is the SNP conferring lactase persistence, which allows adults in certain populations to digest milk, providing a significant nutritional advantage where dairy farming is prevalent. Another compelling case is the sickle cell trait, caused by a single point mutation, which confers resistance to malaria in heterozygous individuals. These examples demonstrate how small genetic changes can dramatically alter fitness in specific environments, driving adaptation and shaping population genetics over time.